Introduction
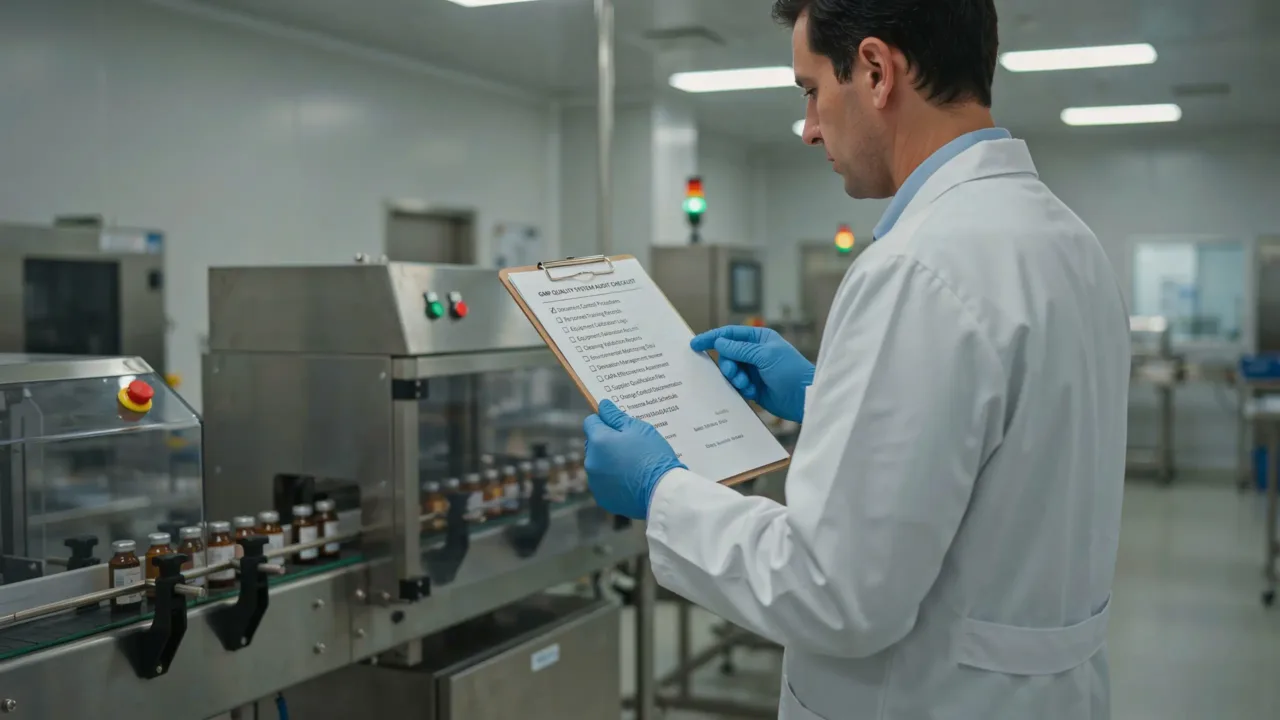
A GMP quality system audit examines the systems, procedures, and processes that ensure pharmaceutical products meet applicable quality and regulatory requirements. It helps organizations identify compliance gaps, evaluate quality management practices, and determine whether corrective actions are effective.
In 2026, pharmaceutical companies need to pay close attention to documentation control, data integrity, supplier qualification, deviation management, and corrective and preventive actions (CAPA). A structured GMP quality system audit checklist helps quality teams review these areas systematically and prepare for regulatory inspections.
This guide covers 15 essential areas to review during a pharmaceutical quality system audit. GxP Cellators provides GMP auditing services to help organizations evaluate their quality systems, identify potential risks, and develop practical improvement plans.
GMP Quality System Audit Checklist: 15 Areas to Review
1. Quality Management System
The Quality Management System (QMS) provides the foundation for maintaining product quality and regulatory compliance. Auditors should evaluate whether the system is documented, implemented, and regularly reviewed.
- Is there a documented and approved quality management system?
- Is the quality manual current and approved?
- Are quality objectives defined, monitored, and reviewed?
- Are responsibilities for maintaining the QMS clearly assigned?
An effective QMS should support consistent processes, risk management, continuous improvement, and compliance with applicable GMP requirements.
2. Management Responsibility
Management plays an important role in maintaining a strong pharmaceutical quality system. Auditors should assess whether senior management provides the necessary resources, oversight, and direction to support quality objectives.
- Is management committed to product quality and regulatory compliance?
- Are roles, responsibilities, and authorities clearly defined?
- Are management reviews conducted at planned intervals?
- Are sufficient resources available to maintain quality operations?
Management involvement is important because unresolved quality issues may indicate weaknesses in the overall quality management system.
3. Documentation Control
Accurate documentation is essential for demonstrating that pharmaceutical activities are performed according to approved procedures. Documentation control ensures that employees use current instructions and that records remain complete and traceable.
- Are documents reviewed, approved, and controlled?
- Are standard operating procedures (SOPs) current?
- Are obsolete documents removed from active use?
- Are records retained according to applicable requirements?
Auditors should also examine document revisions, approval histories, record corrections, and controls over electronic documentation. Poor documentation practices can create compliance gaps and make it difficult to demonstrate that required activities were performed correctly.
4. Training and Competency
Employees must have the knowledge and skills required to perform their assigned responsibilities. A GMP training audit evaluates whether personnel receive appropriate training before performing regulated activities and whether their competency is maintained.
- Are personnel trained on relevant SOPs and GMP requirements?
- Are training records complete and current?
- Is training effectiveness evaluated?
- Are employees retrained when procedures or responsibilities change?
Training records should demonstrate that employees understand the procedures relevant to their roles. Where necessary, organizations should use competency assessments or practical evaluations rather than relying only on attendance records.
5. Deviation Management
Deviation management ensures that departures from approved procedures, specifications, or expected results are documented and investigated. Effective investigations help organizations understand what happened and prevent similar issues from recurring.
- Are deviations documented promptly?
- Are investigations thorough and scientifically justified?
- Are root causes evaluated appropriately?
- Are corrective actions implemented and monitored?
Auditors should review deviation trends, investigation timelines, recurring issues, and the effectiveness of actions taken. Repeated deviations may indicate underlying weaknesses in procedures, training, equipment, or process controls.
6. Change Control
Change control helps organizations assess and manage changes that could affect product quality, validated systems, or regulatory compliance. Changes should be evaluated and approved before implementation, where required by the applicable procedure.
- Are proposed changes documented and controlled?
- Are quality and regulatory impact assessments conducted?
- Are changes reviewed and approved by authorized personnel?
- Are required validation, documentation, and training activities completed?
A GMP quality system audit should examine changes involving equipment, facilities, computerized systems, manufacturing processes, suppliers, and controlled documents. Auditors should also verify that implemented changes are reviewed to confirm the intended outcome.
7. Corrective and Preventive Actions (CAPA)
CAPA is a central component of an effective pharmaceutical quality system. It helps organizations correct identified problems, investigate their causes, and reduce the likelihood of recurrence.
- Are CAPAs documented and tracked?
- Are root causes identified using appropriate investigation methods?
- Are corrective and preventive actions assigned clear responsibilities?
- Is CAPA effectiveness verified and documented?
Auditors should check whether actions address the actual root cause instead of only correcting the immediate problem. Overdue CAPAs, repeated findings, and ineffective actions should receive appropriate attention based on their potential impact on product quality and patient safety.
8. Supplier Management
Supplier management ensures that materials, components, and services obtained from external providers meet established quality requirements. Supplier oversight is particularly important when external organizations can affect product quality or GMP compliance.
- Are suppliers evaluated and qualified before approval?
- Are supplier audits conducted according to risk and established procedures?
- Are quality agreements in place where appropriate?
- Are supplier performance and quality issues monitored?
A supplier audit should consider the supplier’s quality system, relevant documentation, deviation handling, change notification practices, and previous performance. Supplier qualification and requalification should reflect the level of risk associated with the supplied materials or services.
9. Validation and Qualification
Validation and qualification activities provide documented evidence that processes, equipment, utilities, analytical methods, and computerized systems are suitable for their intended purposes.
- Are relevant manufacturing processes validated?
- Are analytical methods validated or verified, as applicable?
- Are equipment and utilities qualified?
- Are computerized systems validated according to their intended use and risk?
Auditors should review validation protocols, acceptance criteria, reports, deviations, and ongoing monitoring arrangements. They should also assess whether changes are evaluated for their potential impact on the validated state.
10. Data Integrity
Data integrity is essential for maintaining reliable records and demonstrating compliance with pharmaceutical regulations. Information should remain accurate, complete, consistent, and traceable throughout its lifecycle.
- Are appropriate audit trails enabled and reviewed for relevant systems?
- Are user access controls defined and maintained?
- Is data protected against unauthorized alteration or deletion?
- Are electronic and paper records reviewed for completeness and accuracy?
Auditors should examine how data is created, processed, reviewed, stored, and retained. They should also evaluate relevant backup arrangements, access permissions, audit trail review practices, and controls over original records.
Where applicable, organizations should consider ALCOA+ principles: data should be attributable, legible, contemporaneous, original, accurate, complete, consistent, enduring, and available.
11. Self-Inspection and Internal Audits
Self-inspections and internal audits help organizations identify quality weaknesses before they develop into more significant compliance problems. Their effectiveness depends on appropriate planning, objective assessment, and timely follow-up.
- Are self-inspections conducted according to an approved schedule?
- Are audits performed by appropriately qualified and independent personnel?
- Are findings documented and assigned appropriate priorities?
- Are corrective actions tracked through completion and effectiveness checks?
Auditors should review the internal audit programme, previous findings, recurring issues, and evidence of continuous improvement. Audit frequency and scope should reflect operational complexity, risk, and previous compliance performance.
12. Complaints and Recalls
Complaint handling and recall management are important for protecting patients and responding to potential product quality issues. Organizations should maintain procedures for investigating complaints and determining whether further action is necessary.
- Are product complaints documented and investigated?
- Are investigations completed within established timelines?
- Are complaint trends reviewed for recurring quality issues?
- Are recall procedures documented and periodically evaluated?
Auditors should also examine escalation procedures, relevant regulatory notifications, traceability arrangements, and communication responsibilities. Complaint investigations should generate appropriate corrective actions when recurring or significant problems are identified.
13. Annual Product Quality Review
Annual Product Quality Review, or the applicable periodic product quality review process, helps organizations evaluate manufacturing consistency and identify trends that may affect product quality.
- Are periodic product quality reviews conducted as required?
- Are deviations, complaints, and rejected batches evaluated?
- Are product quality and process performance trends identified?
- Are appropriate follow-up actions documented?
Reviews should consider relevant information such as changes, CAPA, stability results, and other applicable quality data. Auditors should verify that identified trends are investigated and that actions are assigned and followed up appropriately.
14. Stability Programme
A pharmaceutical stability programme provides information about product quality over time under defined storage conditions. It supports shelf-life decisions and helps identify potential changes in product characteristics.
- Is stability testing conducted according to approved protocols?
- Are stability results reviewed and documented?
- Are adverse trends investigated?
- Are out-of-specification results handled according to approved procedures?
Auditors should review stability protocols, testing schedules, data records, investigations, and reporting arrangements. They should also verify that significant findings are assessed for their potential impact on product quality, shelf life, and regulatory commitments.
15. Management Review and Quality Metrics
Management review helps senior leaders evaluate the overall performance of the pharmaceutical quality system. It provides an opportunity to assess significant risks, recurring issues, resource requirements, and improvement priorities.
- Are management reviews conducted at planned intervals?
- Are relevant quality metrics and trends reviewed?
- Are recurring deviations, CAPAs, complaints, and audit findings discussed?
- Are decisions and follow-up actions documented and tracked?
Useful indicators may include overdue CAPAs, repeat deviations, supplier performance, training completion, complaint trends, and audit findings. Metrics should support meaningful decisions rather than simply report activity.
Although management responsibility and management review are closely related, the audit should assess both leadership accountability and the effectiveness of the formal review process.
How GxP Cellators Supports Quality System Audits
GxP Cellators provides GMP quality system audits for pharmaceutical companies seeking to evaluate their compliance arrangements and strengthen quality processes. Its audit support can help organizations identify weaknesses, review documented evidence, and prioritize improvements based on operational and compliance risks.
Our Quality System Audit Services Include
- Full-scope quality system audits
- Documentation and SOP reviews
- Data integrity audits
- CAPA effectiveness reviews
- GMP supplier audits
- Mock inspections and inspection-readiness assessments
- CAPA verification and follow-up
The audit scope can be tailored to an organization’s facilities, processes, quality risks, and applicable regulatory expectations.
Why Choose GxP Cellators?
Organizations can benefit from a practical approach to identifying compliance gaps and improving quality system performance.
- Industry and regulatory experience: Auditors with relevant regulatory inspection and pharmaceutical quality experience.
- Regulatory coverage: Consideration of applicable FDA, EMA, Health Canada, MHRA, and WHO requirements.
- Risk-based assessment: Audit scope tailored to operational risks and organizational needs.
- Actionable reporting: Findings and recommendations designed to support practical remediation.
- Confidentiality: Appropriate protection of proprietary information and quality documentation.
A well-planned audit should do more than identify deficiencies. It should help the organization understand their causes, establish appropriate priorities, and implement sustainable improvements.
How to Prepare for a GMP Quality System Audit in 2026
Before an audit, organizations should define the scope, identify applicable requirements, and review previous findings. Preparing relevant SOPs, training records, deviation investigations, CAPA records, supplier documentation, validation reports, and data integrity controls can make the process more effective.
It is also useful to assign responsible personnel to each audit area and confirm that outstanding actions are being tracked. However, preparation should focus on the actual implementation of quality procedures rather than merely organizing documents for the auditor.
After the audit, findings should be assessed according to their significance and potential impact. The organization should establish clear action owners, target dates, root-cause investigations, and effectiveness checks where appropriate.
A risk-based approach helps quality teams focus resources on issues that could have the greatest impact on product quality, patient safety, and regulatory compliance.
Frequently Asked Questions
1. What is a GMP quality system audit?
A GMP quality system audit examines the systems and processes used to maintain pharmaceutical quality and regulatory compliance. It covers areas such as documentation, training, deviation management, change control, CAPA, supplier management, validation, and data integrity.
2. How many areas should be reviewed in a GMP quality system audit?
The audit scope depends on the organization’s activities, risks, and applicable requirements. This checklist covers 15 important areas, including the QMS, management responsibility, documentation, training, deviations, change control, CAPA, suppliers, validation, data integrity, self-inspection, complaints, product quality review, stability, and management review.
3. How often should GMP quality system audits be conducted?
Audit frequency should be based on applicable regulatory requirements, internal procedures, risk, previous findings, and changes in operations. Some areas may require more frequent review. Organizations should establish a documented audit programme rather than assume that one fixed frequency is suitable for every system.
4. What are the most important areas in a GMP quality system audit?
Important areas include documentation control, deviation management, CAPA, change control, supplier qualification, validation, data integrity, and management oversight. The final priorities should reflect the organization’s processes, previous findings, and potential risks to product quality.
5. How does GxP Cellators support quality system audits?
GxP Cellators provides GMP quality system audit support, documentation reviews, data integrity audits, CAPA effectiveness reviews, supplier audits, mock inspections, and follow-up activities. The scope can be adapted to the organization’s requirements and quality objectives.
6. What should happen after a GMP quality system audit?
After the audit, findings should be documented, evaluated, and prioritized. The organization should investigate root causes, assign corrective actions, establish target dates, and verify effectiveness where appropriate. Follow-up helps ensure that identified issues are addressed and do not continue to affect quality system performance.
7. How can I contact GxP Cellators for quality system audit support?
You can contact GxP Cellators through the official contact page to discuss your audit requirements, quality system concerns, and potential areas for improvement.
Contact GxP Cellators: https://www.gxpcellators.com/contact/
Contact GxP Cellators
If your organization needs support with GMP quality system audits, documentation reviews, data integrity assessments, or CAPA effectiveness reviews, GxP Cellators can help you evaluate your quality systems and identify practical improvement opportunities.
Visit the contact page to discuss your pharmaceutical quality and GMP auditing requirements.

